Last Updated: September 15, 2026
Cleanroom validation is the documented process of proving that a controlled environment consistently meets its classified particulate, airflow, pressure, and microbial limits, and that the evidence will hold up under audit. Applied Physics has supported contamination control programs since 1992, and the single most common mistake we see is procurement teams treating validation as a commodity purchase. It is not. A validation provider is effectively signing your compliance record for the next 6 to 12 months.
Most cleanroom validation scopes include airborne particulate counts, HEPA filter integrity testing, airflow visualization, pressure differential verification, and environmental monitoring. The provider executes these tests, records results, and issues reports that your quality unit files as objective evidence. If the provider’s methods are weak, your data is weak, and no amount of internal documentation fixes that.
Below, we break down exactly how to choose a cleanroom validation service provider, including the credentials that matter, the red flags that should end a conversation, and a checklist you can take into your next vendor audit.
Watch Out The most expensive validation mistake is not a failed test. It is a passed test that later collapses under regulatory scrutiny because the underlying method, calibration, or documentation was never defensible.
Validation is the evidence-gathering process; certification is the formal attestation that follows it. The two terms get used interchangeably in sales conversations, and that confusion costs procurement teams real money.
A validation provider performs the testing and generates the protocol, raw data, and final report. A certifying body reviews that evidence and issues a certificate stating the room meets its target classification, whether that is an ISO Class 5 zone, a Grade B sterile area, or an equivalent internal designation.
Many providers do both. Some do only one. You need to know which you are buying, because a certificate without a defensible validation package behind it is a document, not proof.
Certification requirements are defined by the standard your facility is classified against, and the provider you hire must be fluent in all of them. For most pharmaceutical, biotech, and sterile compounding operations, that means ISO 14644 for particulate classification and cGMP expectations for documentation, data integrity, and change control (iso.org). Sterile compounding pharmacies also work under USP 797 and USP 800 for hazardous and non-hazardous sterile preparations.
A provider who only knows particulate counting will miss the microbial and pressure elements your auditor will ask about. Ask directly which standards they validate against, and ask them to name the specific clauses they test to.
Auditors look for three things in a cleanroom validation package: traceable calibration, complete raw data, and a protocol that matches the standard you claim to meet. If any of those three is missing, the audit finding is not about your room, it is about your vendor.
ISO 14644-1 governs classification by airborne particulate counts. ISO 14644-3 covers the test methods for filter integrity, airflow, and recovery. cGMP adds the expectation that all of this is documented under controlled procedures with an audit trail. The ISO 14644-1 cleanroom classification standard overview is the reference point your auditor will use, and your provider should be able to walk through it clause by clause.
Accreditation, calibration, and traceability are the three credentials that separate a qualified provider from a vendor with a particle counter and a business card. Accreditation means an independent body has assessed the provider’s methods and competence. Calibration means their instruments were checked against a known reference. Traceability means that reference chains back to a national or international standard.
Ask for calibration certificates on every instrument that will enter your room, not just the headline particle counter. Aerosol photometers used for HEPA filter integrity testing, microbial air samplers, and airflow visualization equipment all need current calibration. A provider who cannot produce certificates on the day of the test is a provider who will cost you a repeat visit.
Pro Tip Request calibration certificates at the quoting stage, not the day of testing. Providers with well-managed instrument programs send them without hesitation. Providers who scramble are telling you something about their documentation culture.
A cleanroom validation protocol should arrive before the technician does, and it should name every test, every acceptance criterion, and every instrument. If the protocol is a generic template with your facility name pasted in, that is a problem. Protocols should reflect your room’s layout, classification, and intended use.
Read the protocol against your own standard operating procedures. The acceptance criteria in the protocol must match the limits your quality unit has committed to. A mismatch discovered during an audit is far more expensive than one caught during review.
Installation Qualification (IQ) documents that equipment and systems were installed as specified. Operational Qualification (OQ) proves they function across their intended operating range. Performance Qualification (PQ) demonstrates they perform reliably under real production conditions.
For a cleanroom, IQ covers the physical build: panels, FFUs, ductwork, and monitoring hardware. OQ covers airflow velocity, filter integrity, and pressure differentials under controlled conditions. PQ covers particulate and microbial performance during actual or simulated operations, often over multiple sampling events.
A provider who skips or shortcuts PQ is delivering a snapshot, not a validated environment.
The clearest red flag is a provider who cannot explain their own test method without deferring to a supervisor. Technicians should know why they are running a test, not just how.
Other warning signs worth taking seriously:
That last point matters more than most buyers realize. Raw data is what makes a report defensible. A summary table with no underlying counts is a claim, not evidence.
Validation pricing tiers generally track scope, not quality, and the cheapest tier almost always excludes something you need. A basic tier typically covers particulate counts and filter integrity. A mid tier adds airflow visualization, pressure differential mapping, and microbial sampling. A comprehensive tier adds IQ/OQ/PQ documentation packages, continuous monitoring integration, and post-validation support.
The cost-benefit question is not “which tier is cheapest” but “which tier prevents a repeat visit.” A failed audit or a re-test costs far more than the difference between tiers, both in direct expense and in production downtime.
| Service Tier | Typical Scope | Best For | Watch For |
|---|---|---|---|
| Basic | Particulate counts, filter integrity | Recertification of stable, low-risk rooms | Excludes microbial and airflow testing |
| Standard | Adds airflow visualization, pressure mapping | Most GMP and ISO classified rooms | Confirm raw data is included |
| Comprehensive | Adds IQ/OQ/PQ packages, monitoring integration | New builds, USP 797/800 suites, semiconductor fabs | Verify protocol is facility-specific |
Most quotes you receive will not be directly comparable, because providers scope differently. Before you open a single proposal, build a line-item scope sheet that forces every bidder onto the same page. At minimum, list:
When you normalize quotes this way, the apparent low bid often stops being the low bid. A provider who excludes microbial sampling or charges a separate mobilization fee for a second visit is not cheaper; they have simply moved cost off the headline number.
The purchase price of a validation visit is only one line in the total cost of ownership. The larger drivers are usually:
A practical rule most quality teams use: budget for the tier that matches your regulatory exposure, then add a contingency for one repeat visit per year. If the comprehensive tier costs less than that contingency, it is usually the better buy.
The right tier is a function of what your room is used for, not how big it is. A small sterile compounding suite under USP 797 carries more documentation and microbial sampling burden than a much larger non-sterile packaging area. Semiconductor and medical device environments have their own classification and contamination control expectations. Map your facility to the standard it must meet, then choose the tier that produces evidence for every clause in that standard. Anything less is a gap your auditor will find.
Key Takeaway Match the tier to your regulatory exposure, not your budget line. A sterile compounding pharmacy under USP 797 needs more than a particulate count, even if the room is small.
Cleanroom validation frequency is set by your standard, your classification, and your change control program. ISO 14644-2 provides guidance on the maximum time interval between classifications, and most facilities fall into a 6-month or 12-month cycle depending on class and risk. Any change to the room, including new equipment, panel modifications, or HVAC work, triggers revalidation regardless of the calendar.
The mistake is treating validation as an isolated event. It should be integrated with your ongoing maintenance program so that filter changes, FFU servicing, and monitoring system calibration feed directly into the next validation cycle. Facilities that separate the two end up re-testing the same problems every cycle.
Applied Physics builds this integration into monitoring deployments through our FDA guidance on process validation for medical devices aligned approach, so validation data and maintenance records live in the same system of record. Our Cleanroom Monitoring System (CRMS) continuously tracks airborne particle counts, differential pressure, temperature, and humidity, which means the next validation cycle starts with real trend data instead of a blank slate.
Run this checklist during the vendor audit, before any contract is signed. The answers tell you more than the proposal document ever will. Treat it as a scoring tool, not a conversation starter: for each item, note whether the provider answered confidently, deflected, or could not answer at all.
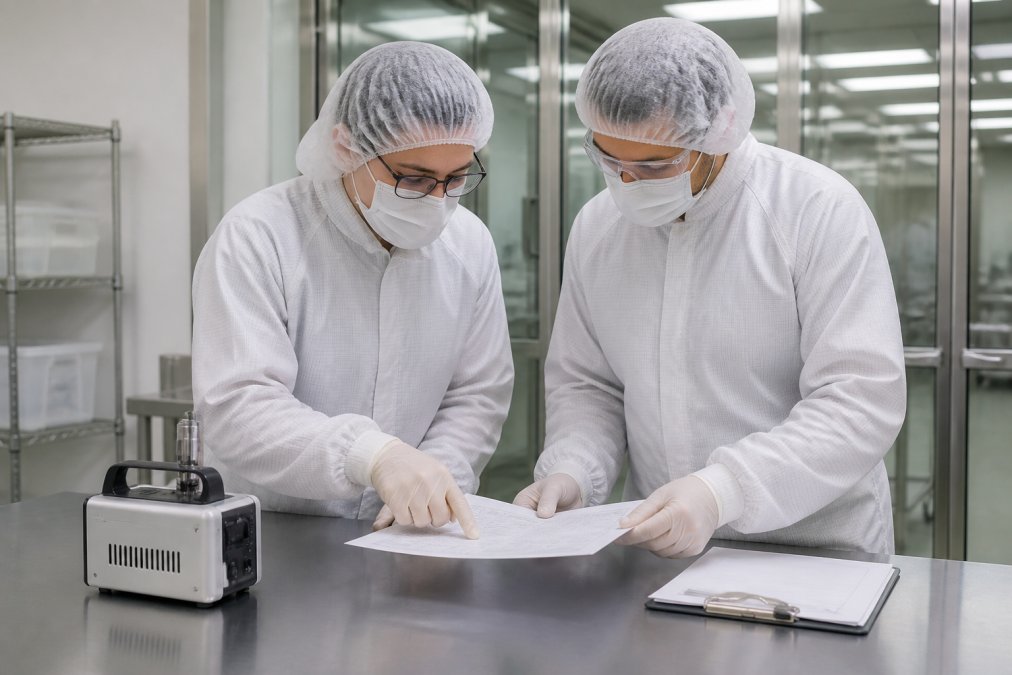
A cleanroom manager and a validation engineer in full gowning reviewing a printed audit checklist on a stainless steel table inside a cleanroom, with a particle counter and clipboard visible
A provider who answers all items confidently is worth a premium. A provider who deflects on three or more is not worth the discount. Use a simple three-point scale for each question:
Add the scores. A total below roughly 70 percent of the maximum is a signal to keep looking, regardless of price. The pattern of zeros matters more than the total: a zero on raw data or calibration is a hard stop, while a zero on turnaround time may be negotiable.
Score the checklist, then overlay the red flags from the previous section. A provider can score well on the scripted questions and still show warning signs in how they operate. Watch for:
That last point matters more than most buyers realize. Raw data is what makes a report defensible. A summary table with no underlying counts is a claim, not evidence.
The audit does not end at the proposal stage. If the provider will perform the work on your site, ask to observe a live test or review a recent completed package. Watching a technician explain their method in real time tells you more than any written answer. A provider who is comfortable being observed is usually a provider whose documentation culture is already strong.
For HEPA filter integrity and airflow work specifically, the tools matter as much as the technician. Applied Physics supplies the Aerosol Photometer BAP-350 for HEPA validation and leak detection, and our LN2 ultrapure and ultrasonic cleanroom foggers for airflow visualization. Facilities that run their own validation internally use these tools alongside third-party testing to cross-check results.
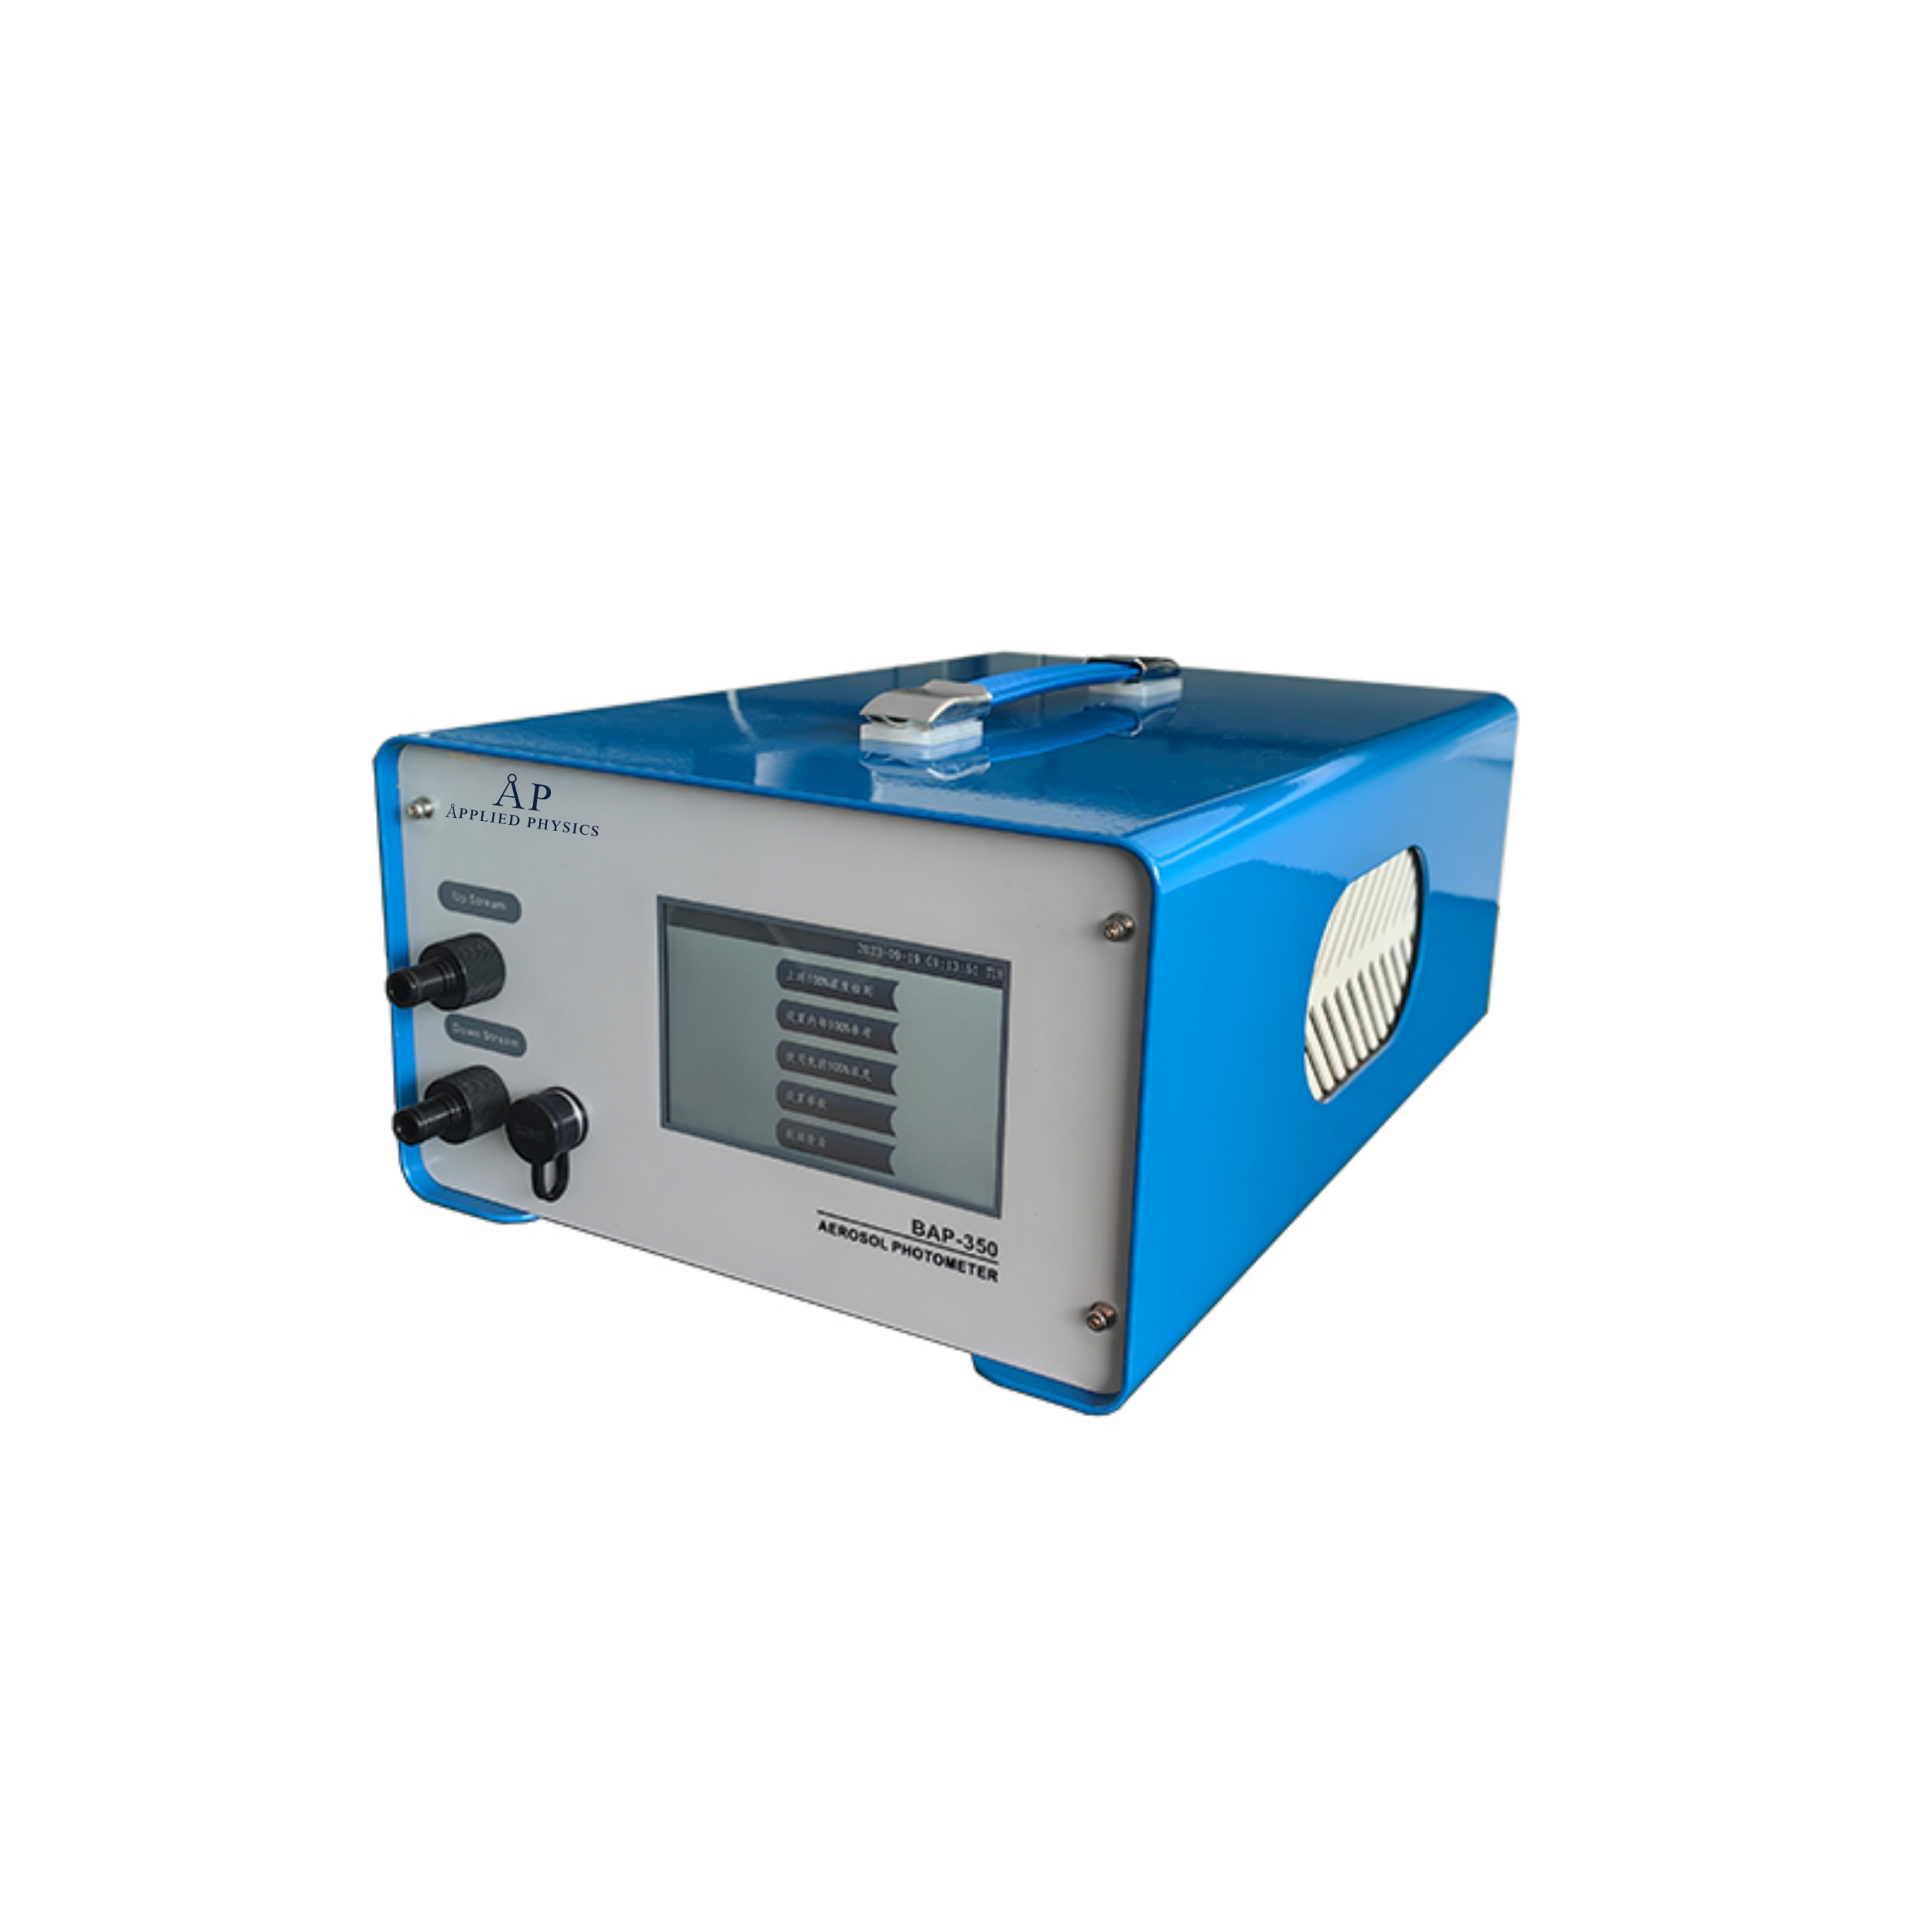
Aerosol Photometer BAP-350
Look for ISO 17025 accreditation for the calibration laboratory, certified cleanroom professionals on staff, and documented experience with your specific regulatory framework: ISO 14644, cGMP, or USP 797/800. Ask for calibration certificates showing traceability to NIST standards. The provider should also carry professional liability insurance and be able to supply references from facilities similar to yours in size and classification.
ISO 14644-2 recommends revalidation every 6 to 12 months for most cleanroom classifications, with high-risk areas like sterile compounding pharmacies often requiring testing every 6 months under USP 797. Semiconductor fabs running sub-10nm processes may validate quarterly. Your cleanroom validation frequency also depends on facility changes, maintenance events, and any excursion that triggers corrective action.
A complete report should contain airborne particulate counts at each sample location, HEPA filter integrity test results, airflow visualization records, pressure differential measurements, and temperature and humidity data. It must include the calibration certificates for every instrument used, the cleanroom validation protocol followed, pass/fail criteria tied to your ISO classification, and the signature of the qualified individual who performed the work.
Validation proves your cleanroom meets its design specification at a point in time, using a defined cleanroom validation protocol with IQ, OQ, and PQ phases. Routine monitoring tracks ongoing performance through continuous particle counting, differential pressure logging, and periodic microbial sampling. Validation is event-driven and documentation-heavy; monitoring is continuous and trend-based. Both are required for audit readiness, and your provider should support both.
Enter your details below to save your shopping cart for later. And, who knows, maybe we will even send you a sweet discount code :)
Allow notifications to get real-time updates about your shopping cart and who knows, you may even receive a sweet discount code 😊