Table of Contents
- What Cleanroom Air Quality Validation Actually Involves
- Pre-Validation Preparation: A Checklist Before You Test
- Step 1: HEPA Filter Integrity Testing
- Step 2: Airflow Velocity and Uniformity Testing
- Step 3: Cleanroom Airflow Visualization (Smoke Testing)
- Step 4: Particle Count Testing and Pressure Differential Monitoring
- Step 5: Cleanroom Validation Frequency and Ongoing Monitoring
- Troubleshooting Common Validation Failures
- Frequently Asked Questions
Last Updated: September 13, 2026
What Cleanroom Air Quality Validation Actually Involves
Cleanroom air quality validation is the documented process of proving that a controlled environment consistently meets its classified cleanliness level for airborne particles, airflow, and pressure relationships. Validation is no longer a one-time event you schedule and forget. It’s a lifecycle obligation, and the facilities that treat it that way pass audits with far less friction.
The core insight most guides bury: validation doesn’t test whether your cleanroom is clean. It tests whether your cleanroom can stay clean under real operating conditions, and whether you can prove that with data. That distinction shapes every test below.
The Three Occupancy States: As-Built, At-Rest, Operational
Every validation runs against three distinct states, defined by ISO 14644 (iso.org). As-built is the empty room with systems running but no equipment or people. At-rest adds installed equipment, still no personnel. Operational is the room doing actual work, with staff and processes active.
Most facilities certify at-rest and then discover their operational particle counts fail. That gap is where contamination control programs quietly fall apart.
ISO 14644 and GMP: Which Standard Applies to Your Facility
ISO 14644 governs cleanroom classification and test methods internationally. GMP requirements, enforced by the FDA for pharmaceutical and sterile compounding operations, layer additional expectations on top: documented protocols, calibration records, and data integrity controls (the FDA).
If you make sterile drug products, you answer to both. If you fabricate semiconductors, ISO 14644 plus your customer specifications typically govern. Identify which framework applies before you write a single test protocol, because the acceptance criteria differ.
Pre-Validation Preparation: A Checklist Before You Test
The most common validation failure isn’t a dirty room. It’s arriving on test day with uncalibrated instruments, incomplete documentation, or HVAC problems you could have caught a week earlier. Preparation is where you win or lose the schedule.
Run this checklist before any test:
- All instruments calibrated within their valid window, with certificates on file
- HVAC system balanced and running at design air change rate for at least 24 hours
- All filters installed, sealed, and documented with serial numbers
- Room surfaces cleaned per your contamination control procedure
- Validation protocol approved and signed by QA
- Supply diffusers and returns verified unobstructed
- Personnel trained on gowning and test procedures
- Baseline environmental monitoring data reviewed
HVAC Performance Checks and Documentation Review
Before particle testing, confirm your HVAC system delivers the design air change rate and that supply diffusers produce uniform air distribution. Review as-built drawings against the current room configuration. If the room was modified, your drawings are probably stale, and stale drawings invalidate your baseline.
Documentation review sounds bureaucratic. In practice, it’s the step that prevents a re-test next month.
Step 1: HEPA Filter Integrity Testing
HEPA filter integrity testing is the first and most consequential test in any validation sequence. A single pinhole leak in a filter or its gasket can compromise an entire classified area, and particle counts alone often miss it because the leak is localized.
The test verifies that the filter and its sealing frame prevent unfiltered air from bypassing the filter media. You test every HEPA and ULPA filter, plus the frames and gaskets around them.
Aerosol Photometer Method for Leak Detection
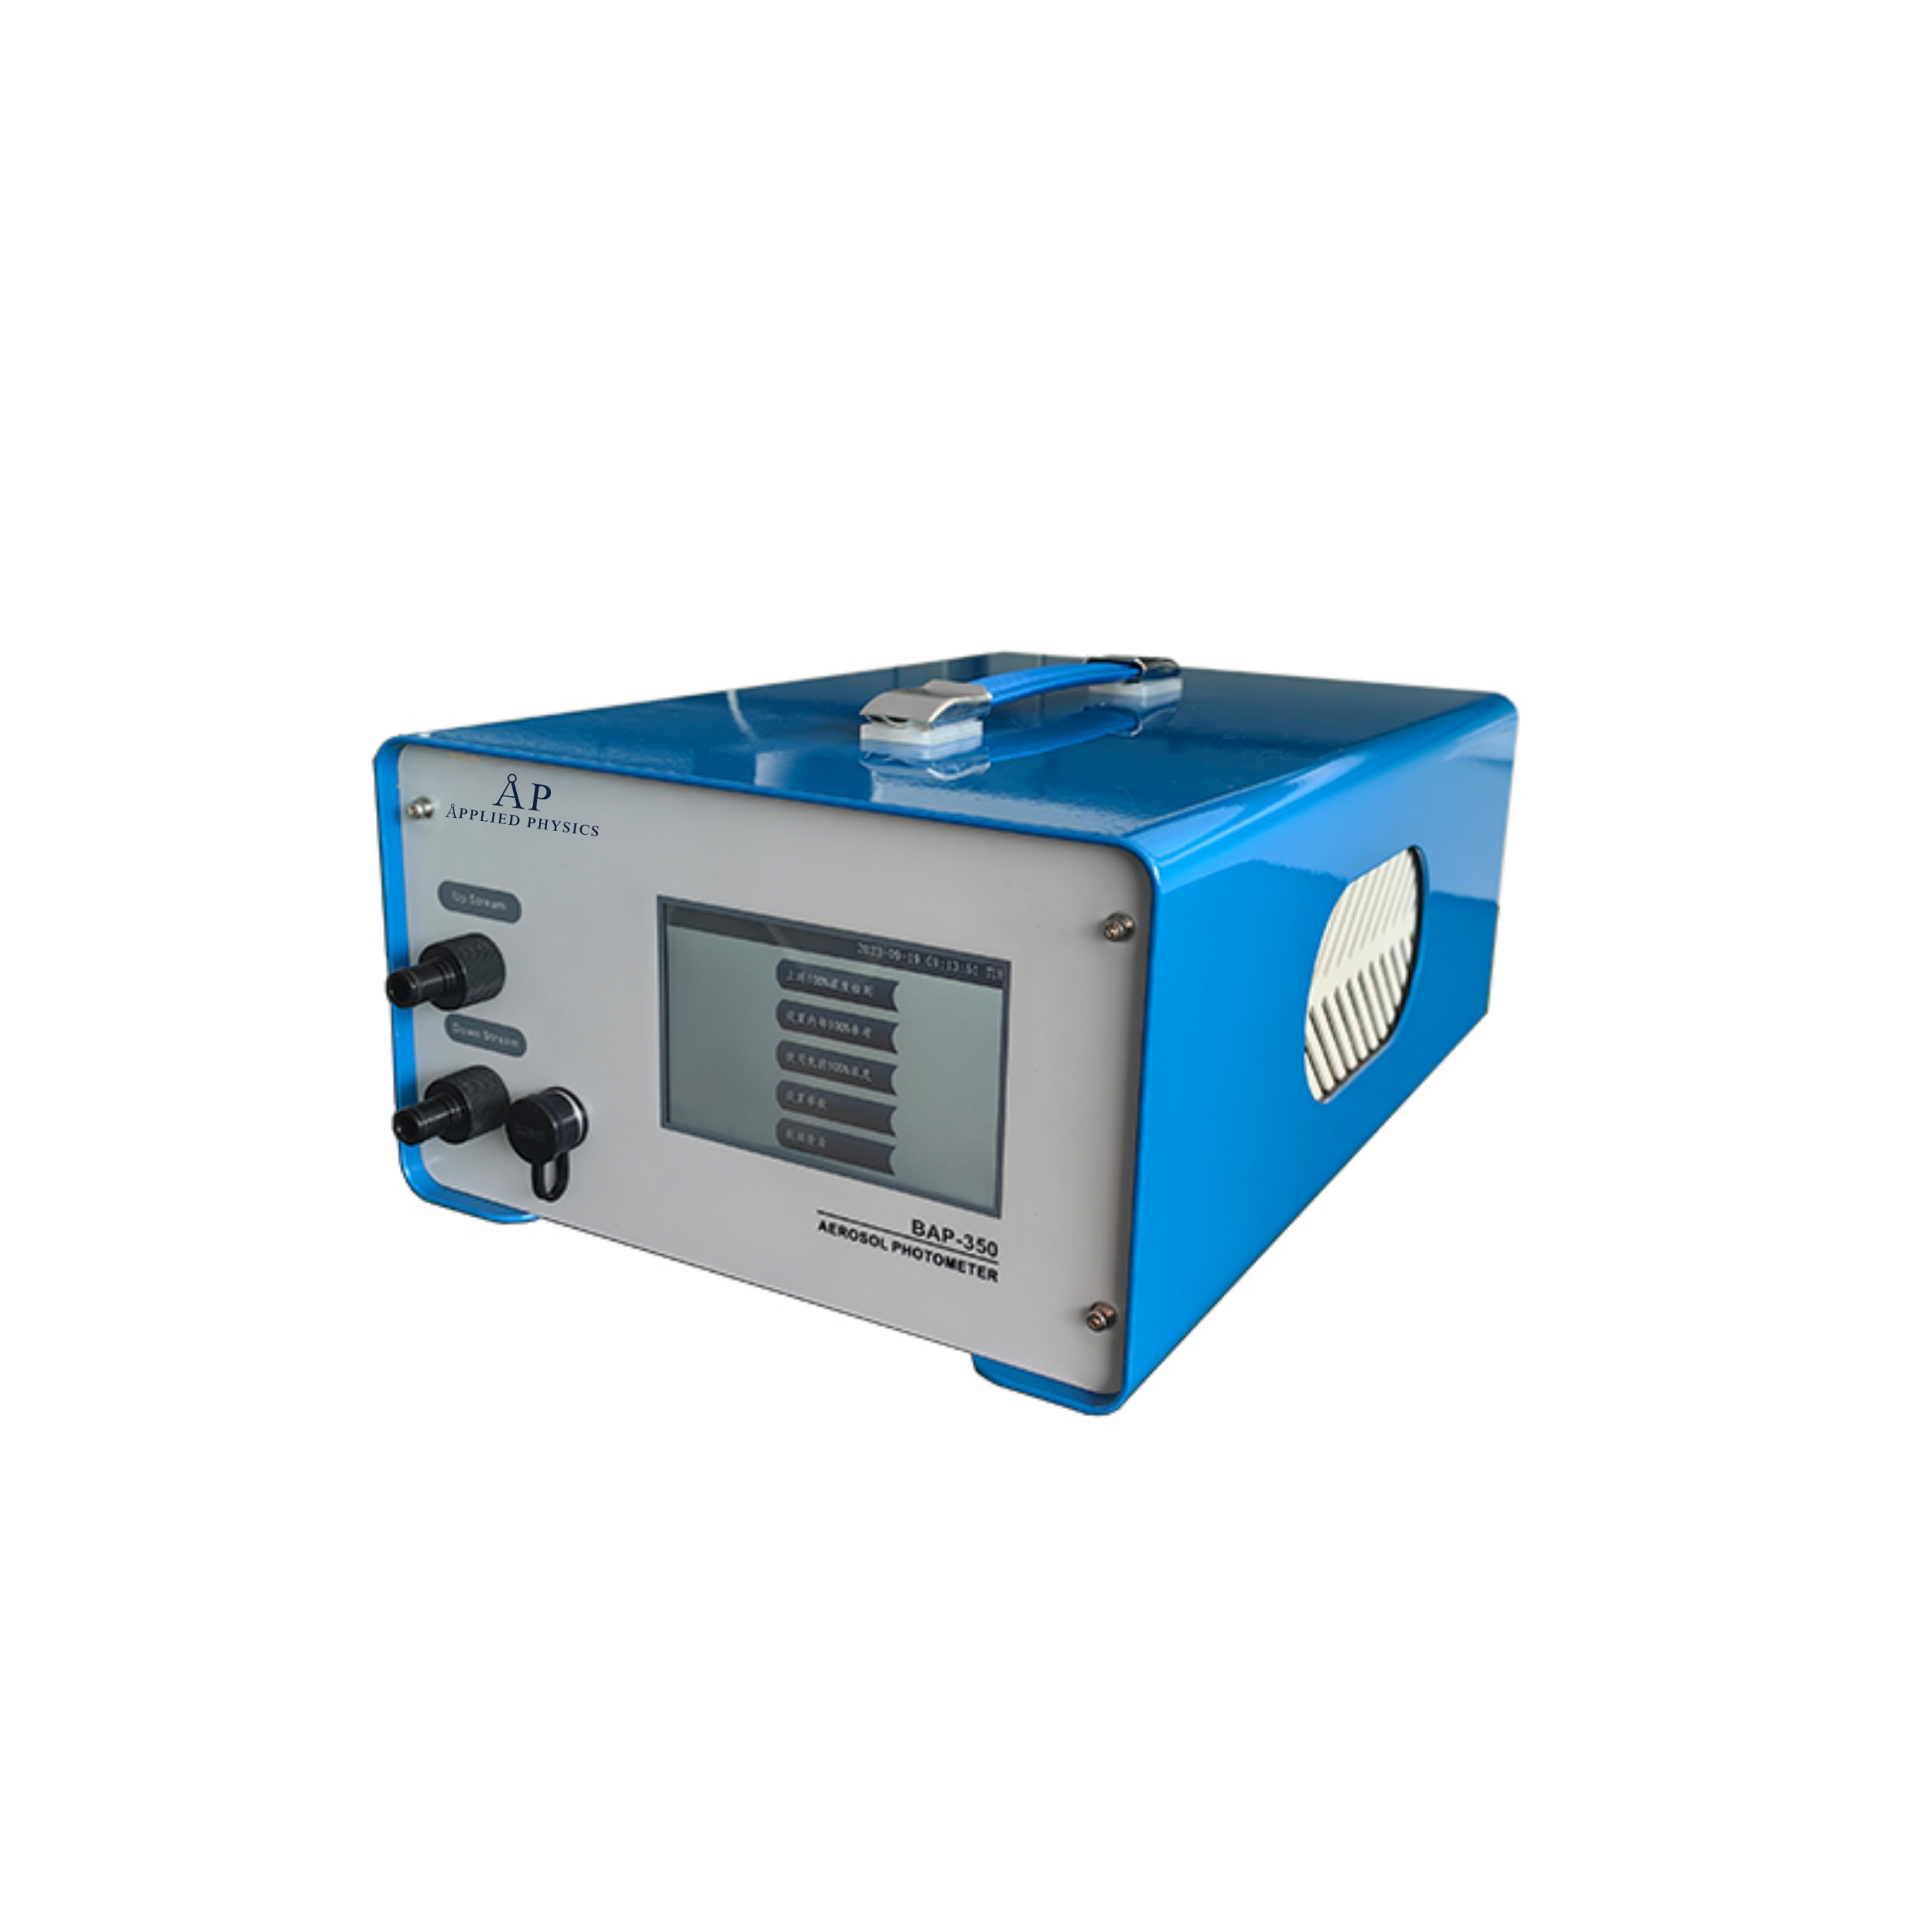
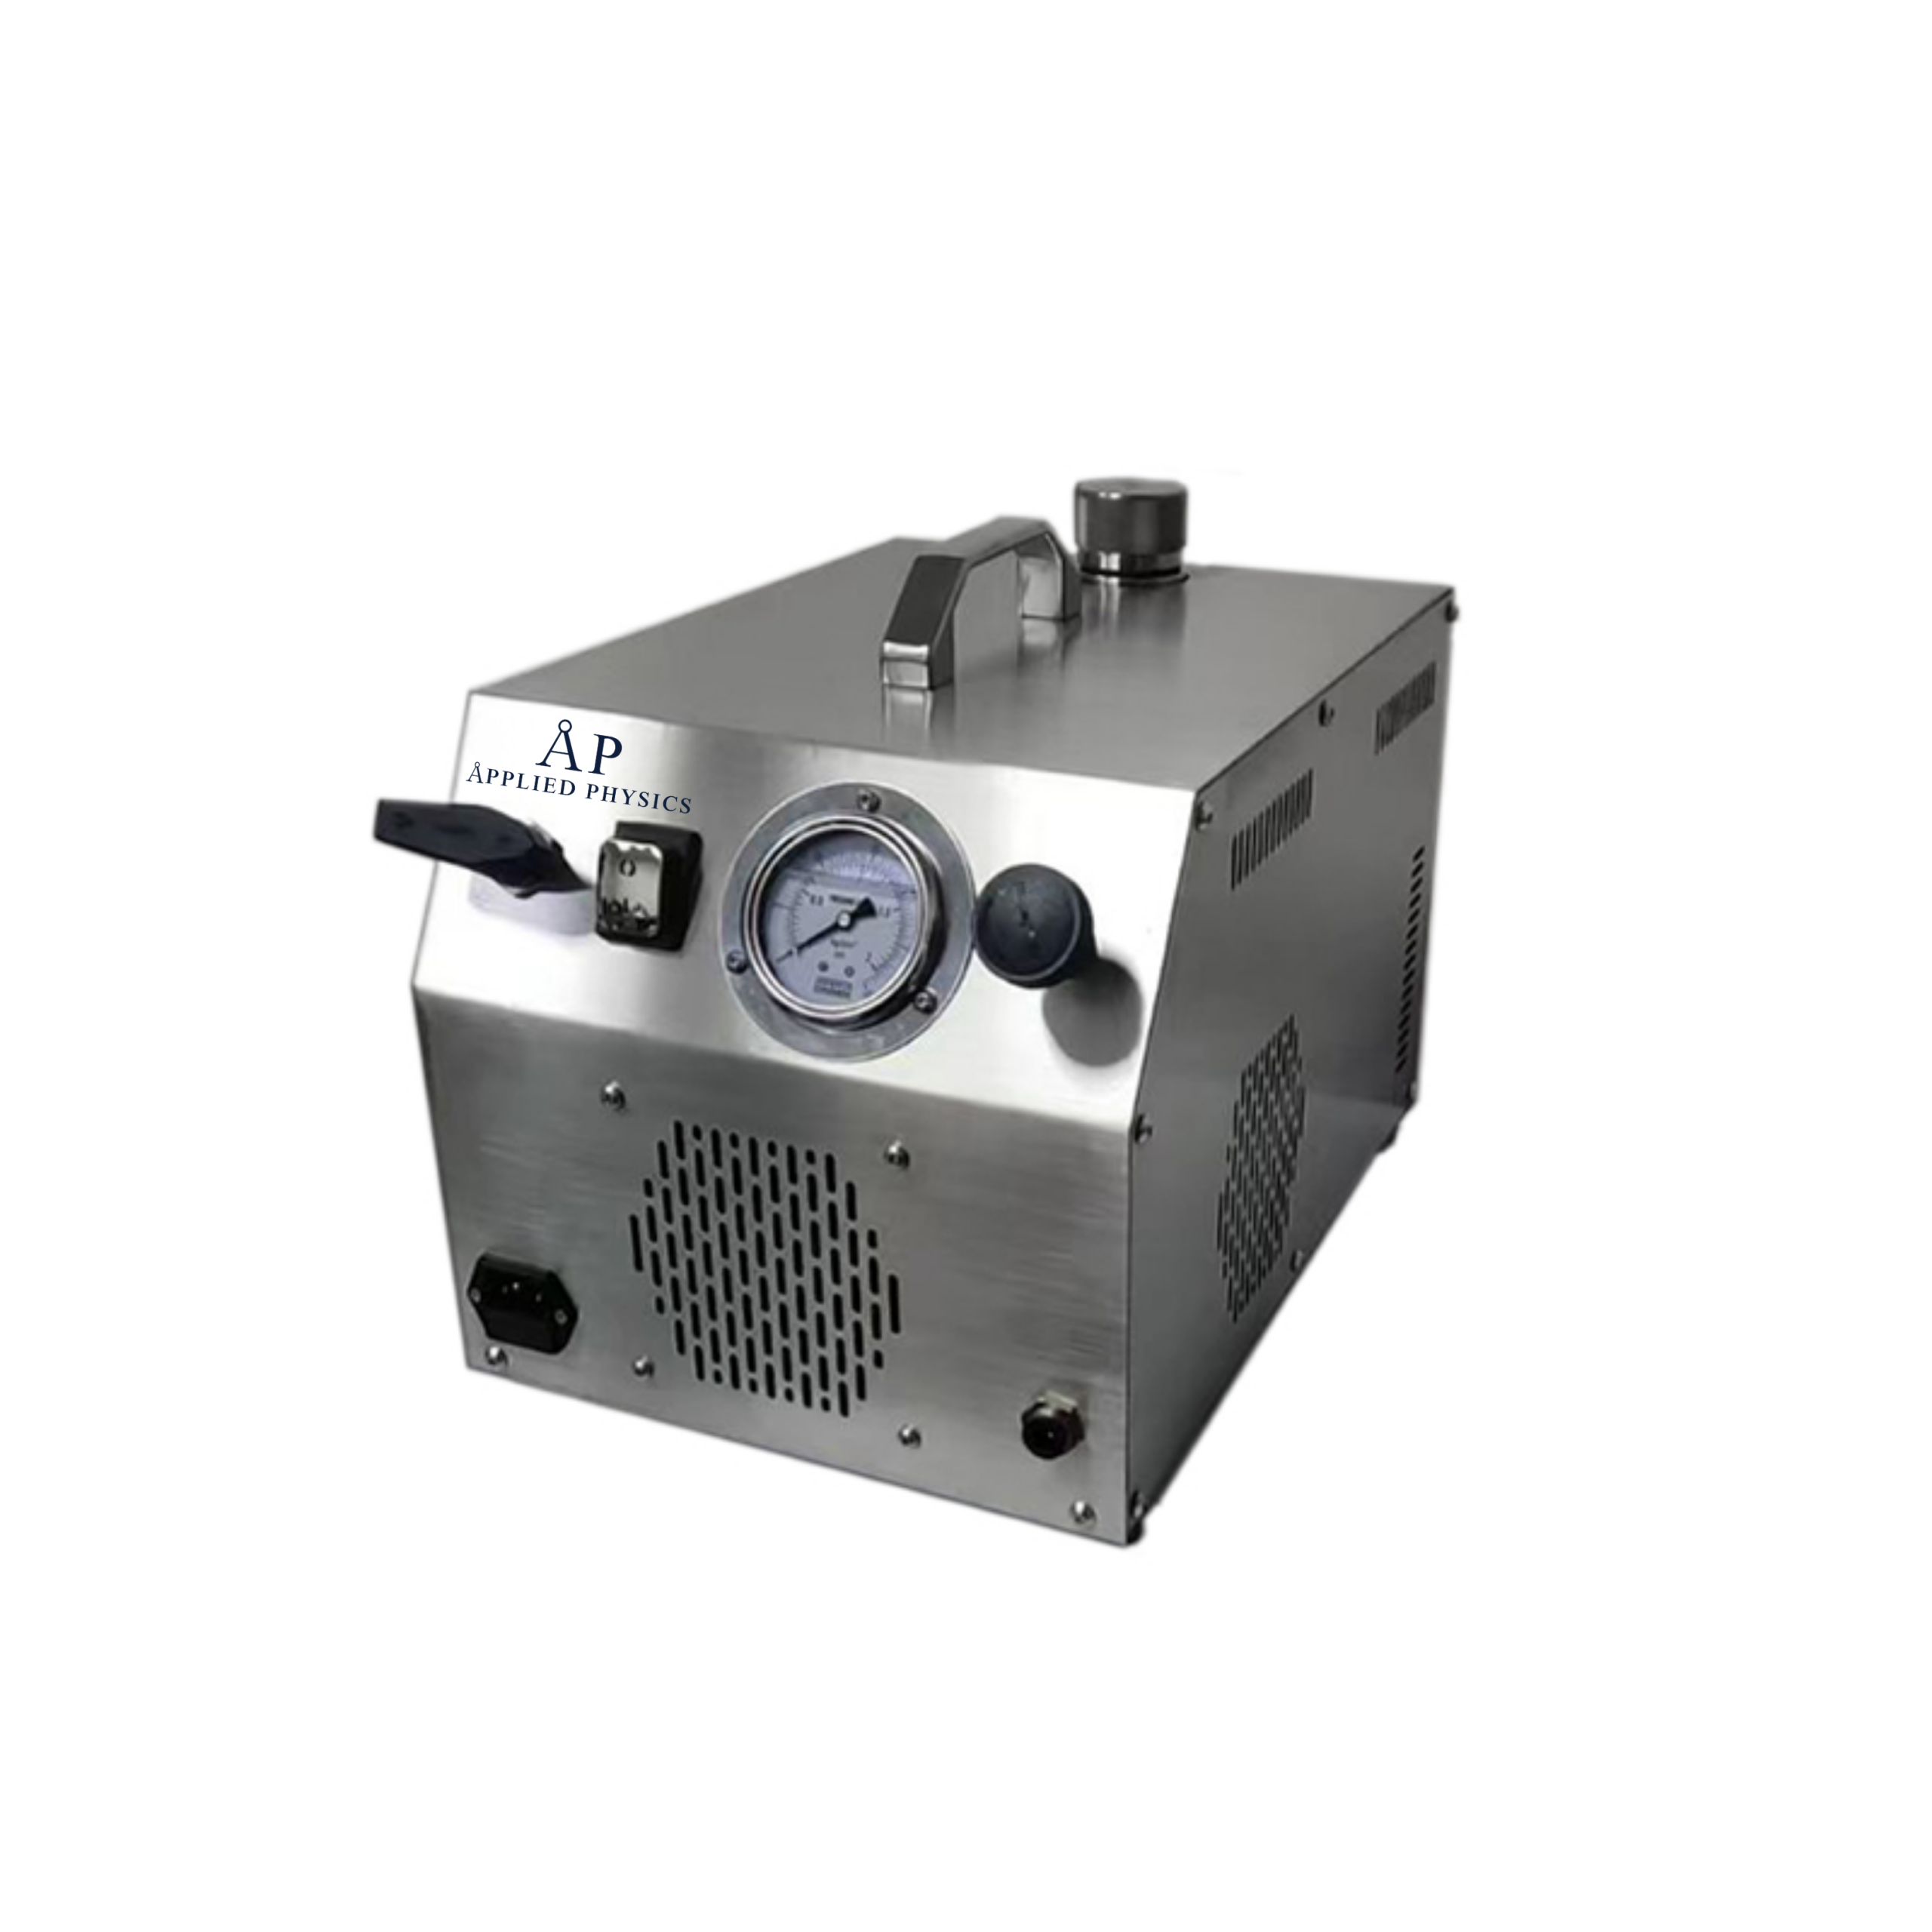
The standard method introduces a challenge aerosol upstream of the filter, then scans the downstream face with a photometer. A common setup pairs an aerosol generator with a photometer: the generator produces a controlled challenge concentration, the photometer measures penetration.
Applied Physics supplies both ends of that workflow. The Aerosol Photometer BAP-350 handles HEPA validation with a compact form factor and a 5-inch color touch screen that speeds up scanning and data capture. For aerosol generation, the Aerosol Generator-Cold Type BAG-6D covers pharmaceutical filter leak detection across clean rooms, laminar flow tables, biosafety cabinets, glove boxes, and HVAC systems.
A common mistake is scanning too fast. Move the probe at a controlled rate, overlap each pass, and pay extra attention to filter frame seals and gasket corners. That’s where leaks hide.
Testing with an uncalibrated photometer is worse than not testing at all. It produces a pass result you can’t defend in an audit, and if a real leak exists, you’ll discover it during a regulatory inspection instead of on your own schedule.
Step 2: Airflow Velocity and Uniformity Testing
Airflow velocity testing confirms that air moves through the cleanroom at the rate your classification requires. For unidirectional flow areas, you measure velocity at the filter face; for non-unidirectional rooms, you calculate air change rate from supply volume.
Use a calibrated anemometer and take readings on a defined grid across each supply diffuser. Record every reading, not an average. Uniformity matters as much as the mean value: a diffuser averaging the right velocity but with dead spots will fail operational particle testing.
The instrument detail that trips people up: anemometers drift, and a five percent calibration error changes your acceptance decision. Verify calibration status the morning of the test, not the week before.
Step 3: Cleanroom Airflow Visualization (Smoke Testing)
Cleanroom airflow visualization uses a visible fog or smoke tracer to reveal how air actually moves through the space. It exposes turbulence, dead zones, and reverse flow that velocity measurements alone cannot show.
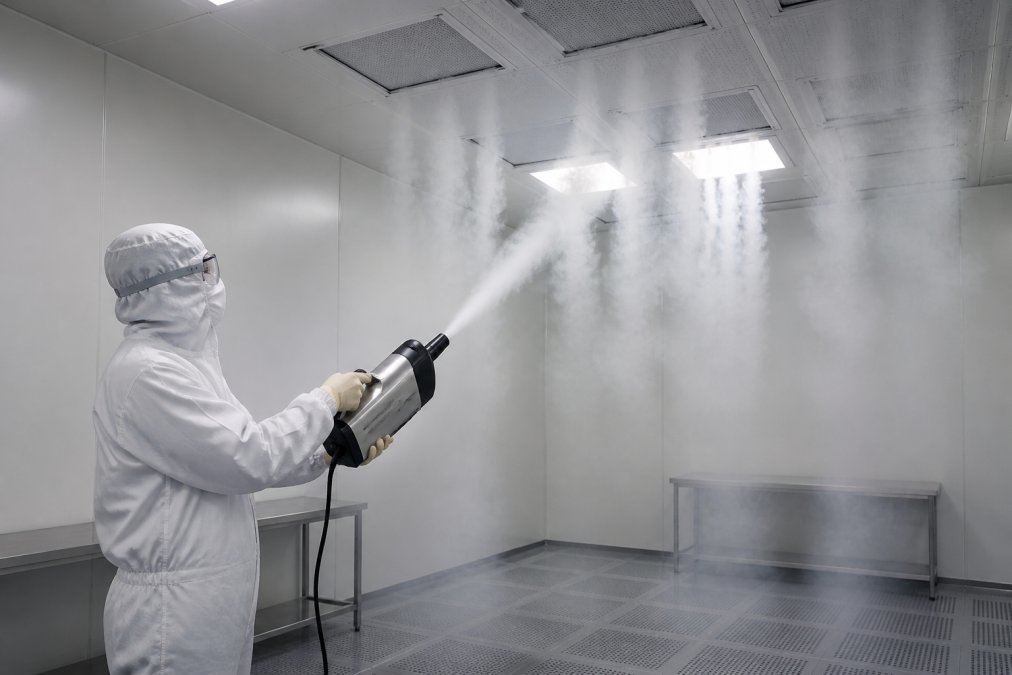
This is the step where most facilities learn something uncomfortable. Velocity readings look fine, particle counts pass at-rest, and then the smoke shows air rolling back toward a critical zone because a piece of equipment disrupted the laminar flow.
Applied Physics offers both LN2 ultrapure and ultrasonic cleanroom foggers. The practical difference matters: a fogger that produces non-neutral buoyancy sends smoke that rises or sinks on its own, and you end up reading the fogger’s behavior instead of the room’s airflow. Neutral-output fog travels with the air stream, which is the entire point.
Film your smoke studies. Reviewing footage frame by frame catches recirculation eddies that are nearly invisible in real time, and the recording doubles as compliance documentation for your validation report.
Step 4: Particle Count Testing and Pressure Differential Monitoring
Particle count testing measures airborne particulate matter at the sizes your classification specifies, typically 0.5 µm and 5.0 µm under ISO 14644 (iso.org). Sample at defined locations, at the required sample volume, in both at-rest and operational states.
Pressure differential monitoring runs alongside it. Your cleanroom must maintain a positive pressure cascade relative to adjacent, less clean spaces, and that differential must hold continuously, not just on test day. Install continuous monitoring rather than relying on spot readings. Maintaining this controlled environment requires a broader commitment to industrial hygiene monitoring to ensure that airborne contaminants remain within acceptable limits throughout the entire facility.
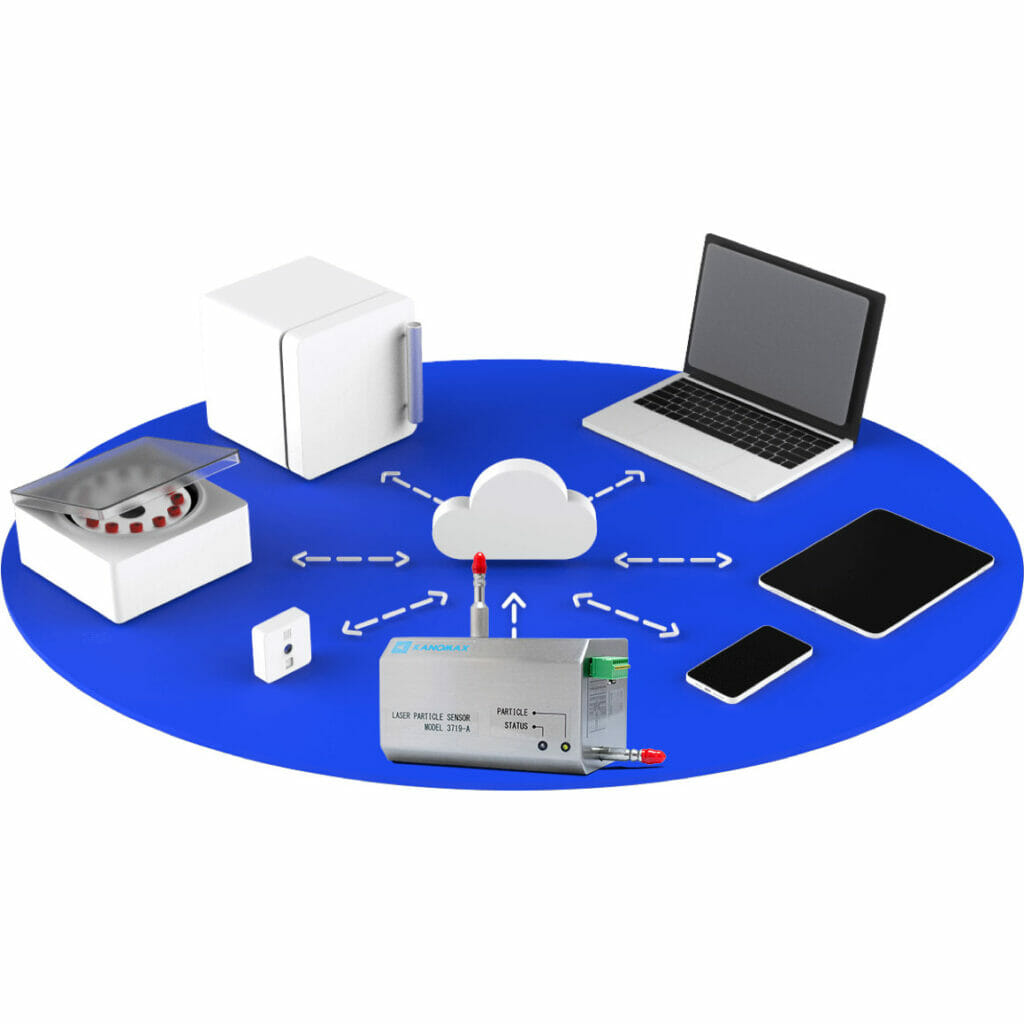
The Cleanroom Monitoring System – Model CRMS handles continuous particle and environmental monitoring, which converts your validation from a snapshot into a live compliance record. For facilities moving to tighter nodes, Applied Physics provides calibration wafer standards and particle metrology.
A representative test sequence looks like this:
| Step | Test | Typical Duration | Frequency |
|---|---|---|---|
| 1 | HEPA filter integrity | 1-2 hours per filter bank | On install, after change |
| 2 | Airflow velocity and uniformity | 2-4 hours | Annually, after modification |
| 3 | Airflow visualization | 2-3 hours | Annually, after layout change |
| 4 | Particle count | 3-6 hours | Quarterly to annually |
| 5 | Pressure differential | Continuous | Monitored continuously |
Step 5: Cleanroom Validation Frequency and Ongoing Monitoring
Cleanroom validation frequency is not a single calendar rule. It is the product of three inputs: your ISO 14644 classification, your regulatory framework, and the rate of change in the room itself. ISO 14644-2 sets the re-qualification framework, and it explicitly ties the monitoring interval to risk rather than to a fixed clock. GMP environments, enforced by the FDA for pharmaceutical and sterile compounding operations, layer a stricter expectation on top: continuous environmental monitoring plus periodic requalification, with the interval justified in writing.
The baseline schedule most facilities land on looks like this:
- Full requalification: annually for most classified spaces, though ISO 14644-2 permits extending the interval when a documented risk assessment and a clean monitoring history support it.
- Particle counts: quarterly for ISO 5 through ISO 7, semi-annually or annually for less critical classifications.
- HEPA filter integrity: on installation, after any filter or gasket change, and at requalification.
- Pressure differential: continuous, not periodic.
- Recovery time: at requalification and after any change to air change rate or layout.
The Change Triggers That Reset Your Baseline
The calendar is the easy part. The harder discipline is recognizing that certain events invalidate part of your validation baseline regardless of where you sit in the annual cycle. A partial requalification is triggered by:
- A HEPA or ULPA filter change, including a like-for-like swap
- Any modification to supply diffusers, returns, or exhaust balance
- A layout change that moves equipment into or out of a critical zone
- A change in occupancy, gowning protocol, or process that alters particle generation
- A reclassification of the space, upward or downward
- A failed monitoring excursion that exceeds your alert or action limits
Treat each of these as a change-control event. The validation question is not “when did we last test?” but “what changed since the last test, and does the existing data still describe the room?”
Why Continuous Monitoring Changes the Frequency Conversation
Spot-checking once a quarter tells you what the room did on four days out of ninety. Continuous monitoring tells you what it did every day, and that distinction is reshaping how facilities justify their requalification intervals. A continuous particle and environmental monitoring system, such as the Cleanroom Monitoring System – Model CRMS, generates the trend data that supports a risk-based interval extension under ISO 14644-2 and produces the audit trail GMP inspectors expect.
The practical trade-off: continuous monitoring carries a higher upfront instrument cost than a handheld particle counter, but it converts your validation from a snapshot into a live compliance record. When an excursion happens at 2 a.m., you have data. When an auditor asks how you know the room held its classification between quarterly tests, you have an answer that a spot-check schedule cannot provide.
Validation frequency is a change-management question, not a calendar question. Every modification to the room, the HVAC, or the process resets part of your baseline, and continuous monitoring is what lets you defend the interval between full requalifications.
Troubleshooting Common Validation Failures
Most guides stop at listing the tests. The harder question, and the one facility managers actually lose sleep over, is what to do when a test fails. This is the diagnostic framework we walk clients through, organized by the test that failed and the most probable root cause.
When HEPA Filter Integrity Testing Fails
A failed scan almost never means the filter media is defective. Work through the causes in this order:
- Gasket and frame seals. The seal between the filter and its housing is the most common leak path. Re-scan the full perimeter before touching the media.
- Filter seating. A filter that is not fully seated in its track leaves a bypass gap that a photometer will catch at the frame.
- Media damage from handling. Pinholes from installation or from contact with tools show up as localized penetration on the downstream face.
- Challenge aerosol problems. If the upstream concentration is unstable or below the required level, the photometer reads low and you get a false pass or an erratic result. Verify the generator output before you condemn the filter.
Repair the gasket rather than patching the media. A patched filter is a temporary fix that reappears at the next scan, and auditors treat patch records as a red flag.
When Particle Counts Pass at-Rest but Fail Operational
This is the single most common operational failure, and it almost always points to airflow disruption rather than a filtration problem. The diagnostic sequence:
- Re-run the airflow visualization study with the room in its operational configuration, including equipment and personnel in position.
- Check whether a piece of equipment has been moved into a critical zone and is breaking unidirectional flow.
- Verify that gowning procedures match what was validated. A change in garment type or donning sequence changes particle generation.
- Confirm that the room is actually at its design air change rate. A supply volume below design will pass at-rest and fail under load.
If the smoke study shows clean, unobstructed flow and the room still fails, the problem is usually generation rate, not removal rate. Look at the process itself.
When Pressure Differentials Drift
A collapsing pressure cascade is a mechanical problem, and the causes are finite:
- Door seals and pass-throughs. A single worn gasket on a high-traffic door can erase your differential. Inspect seals on the doors that open most often.
- Exhaust balance. A change in exhaust volume, from a new fume hood or a rebalanced system, shifts the room’s pressure relationship.
- HVAC damper drift. Dampers move over time. Verify supply and return volumes against the as-built balance report.
- Filter loading. As filters load, resistance rises and supply volume falls. A differential that held last year may not hold this year without a fan adjustment.
When Recovery Time Testing Fails
Recovery time is the test that exposes an underperforming HVAC system. If the room takes too long to return to its classified state after a contamination event, the cause is almost always air change rate below design. Verify supply volume before assuming the filters are at fault. If supply volume is correct and recovery still fails, check for dead zones or short-circuiting revealed by the smoke study.
The Cost of Skipping the Diagnosis
Each of these failures costs a re-test, and a re-test costs a production day in most facilities. The pattern that separates facilities with clean audit histories from those with recurring findings is not better equipment. It is a pre-validation checklist that catches the mechanical issues early and a smoke study run before the particle count, not after it fails.
Do not treat a failed test as a filter problem by default. The majority of failures trace to airflow, sealing, or balance issues that a diagnostic sequence identifies in an hour, while a filter replacement costs a day and often does not fix the underlying cause.
Frequently Asked Questions
What is ISO 14644 cleanroom validation?
ISO 14644 is the international standard that defines cleanroom classification by airborne particle concentration. Validation under this standard involves testing HEPA filter integrity, airflow velocity and uniformity, particle counts at the specified class limit, and pressure differentials. The results determine whether your cleanroom meets its target ISO class, whether that is ISO 5 for sterile compounding or ISO 3 for semiconductor fabrication. Testing must occur in as-built, at-rest, and operational states.
How often should cleanroom air quality be validated?
ISO 14644-2 recommends a maximum testing interval of 6 months for particle count and 12 months for other tests in cleanrooms in continuous operation. High-risk facilities or those with recent modifications may need quarterly testing. After any HVAC change, filter replacement, or room reconfiguration, full revalidation is required regardless of the calendar schedule.
What equipment is needed for cleanroom air quality testing?
You need a particle counter for ISO classification, an aerosol photometer and aerosol generator for HEPA filter integrity testing, an anemometer for airflow velocity measurements, a differential pressure gauge, and a fogger for airflow visualization. For pharmaceutical and semiconductor applications, calibration certificates traceable to NIST standards are required for all instruments. Data logging capability is also important for compliance documentation.
What causes a cleanroom to fail validation?
Common failure points include HEPA filter leaks at gaskets or frames, insufficient air change rates, incorrect pressure differentials between adjacent zones, and turbulent airflow caused by misplaced equipment or supply diffusers. Particle count failures often trace back to inadequate gowning procedures or residual contamination from maintenance work. Reviewing airflow visualization results first usually identifies the root cause faster than re-running particle counts.